Arc Research

Few-shot regulatory sequence design with a genomic language model
BioRxiv
Hie Lab

Few-shot regulatory sequence design with a genomic language model

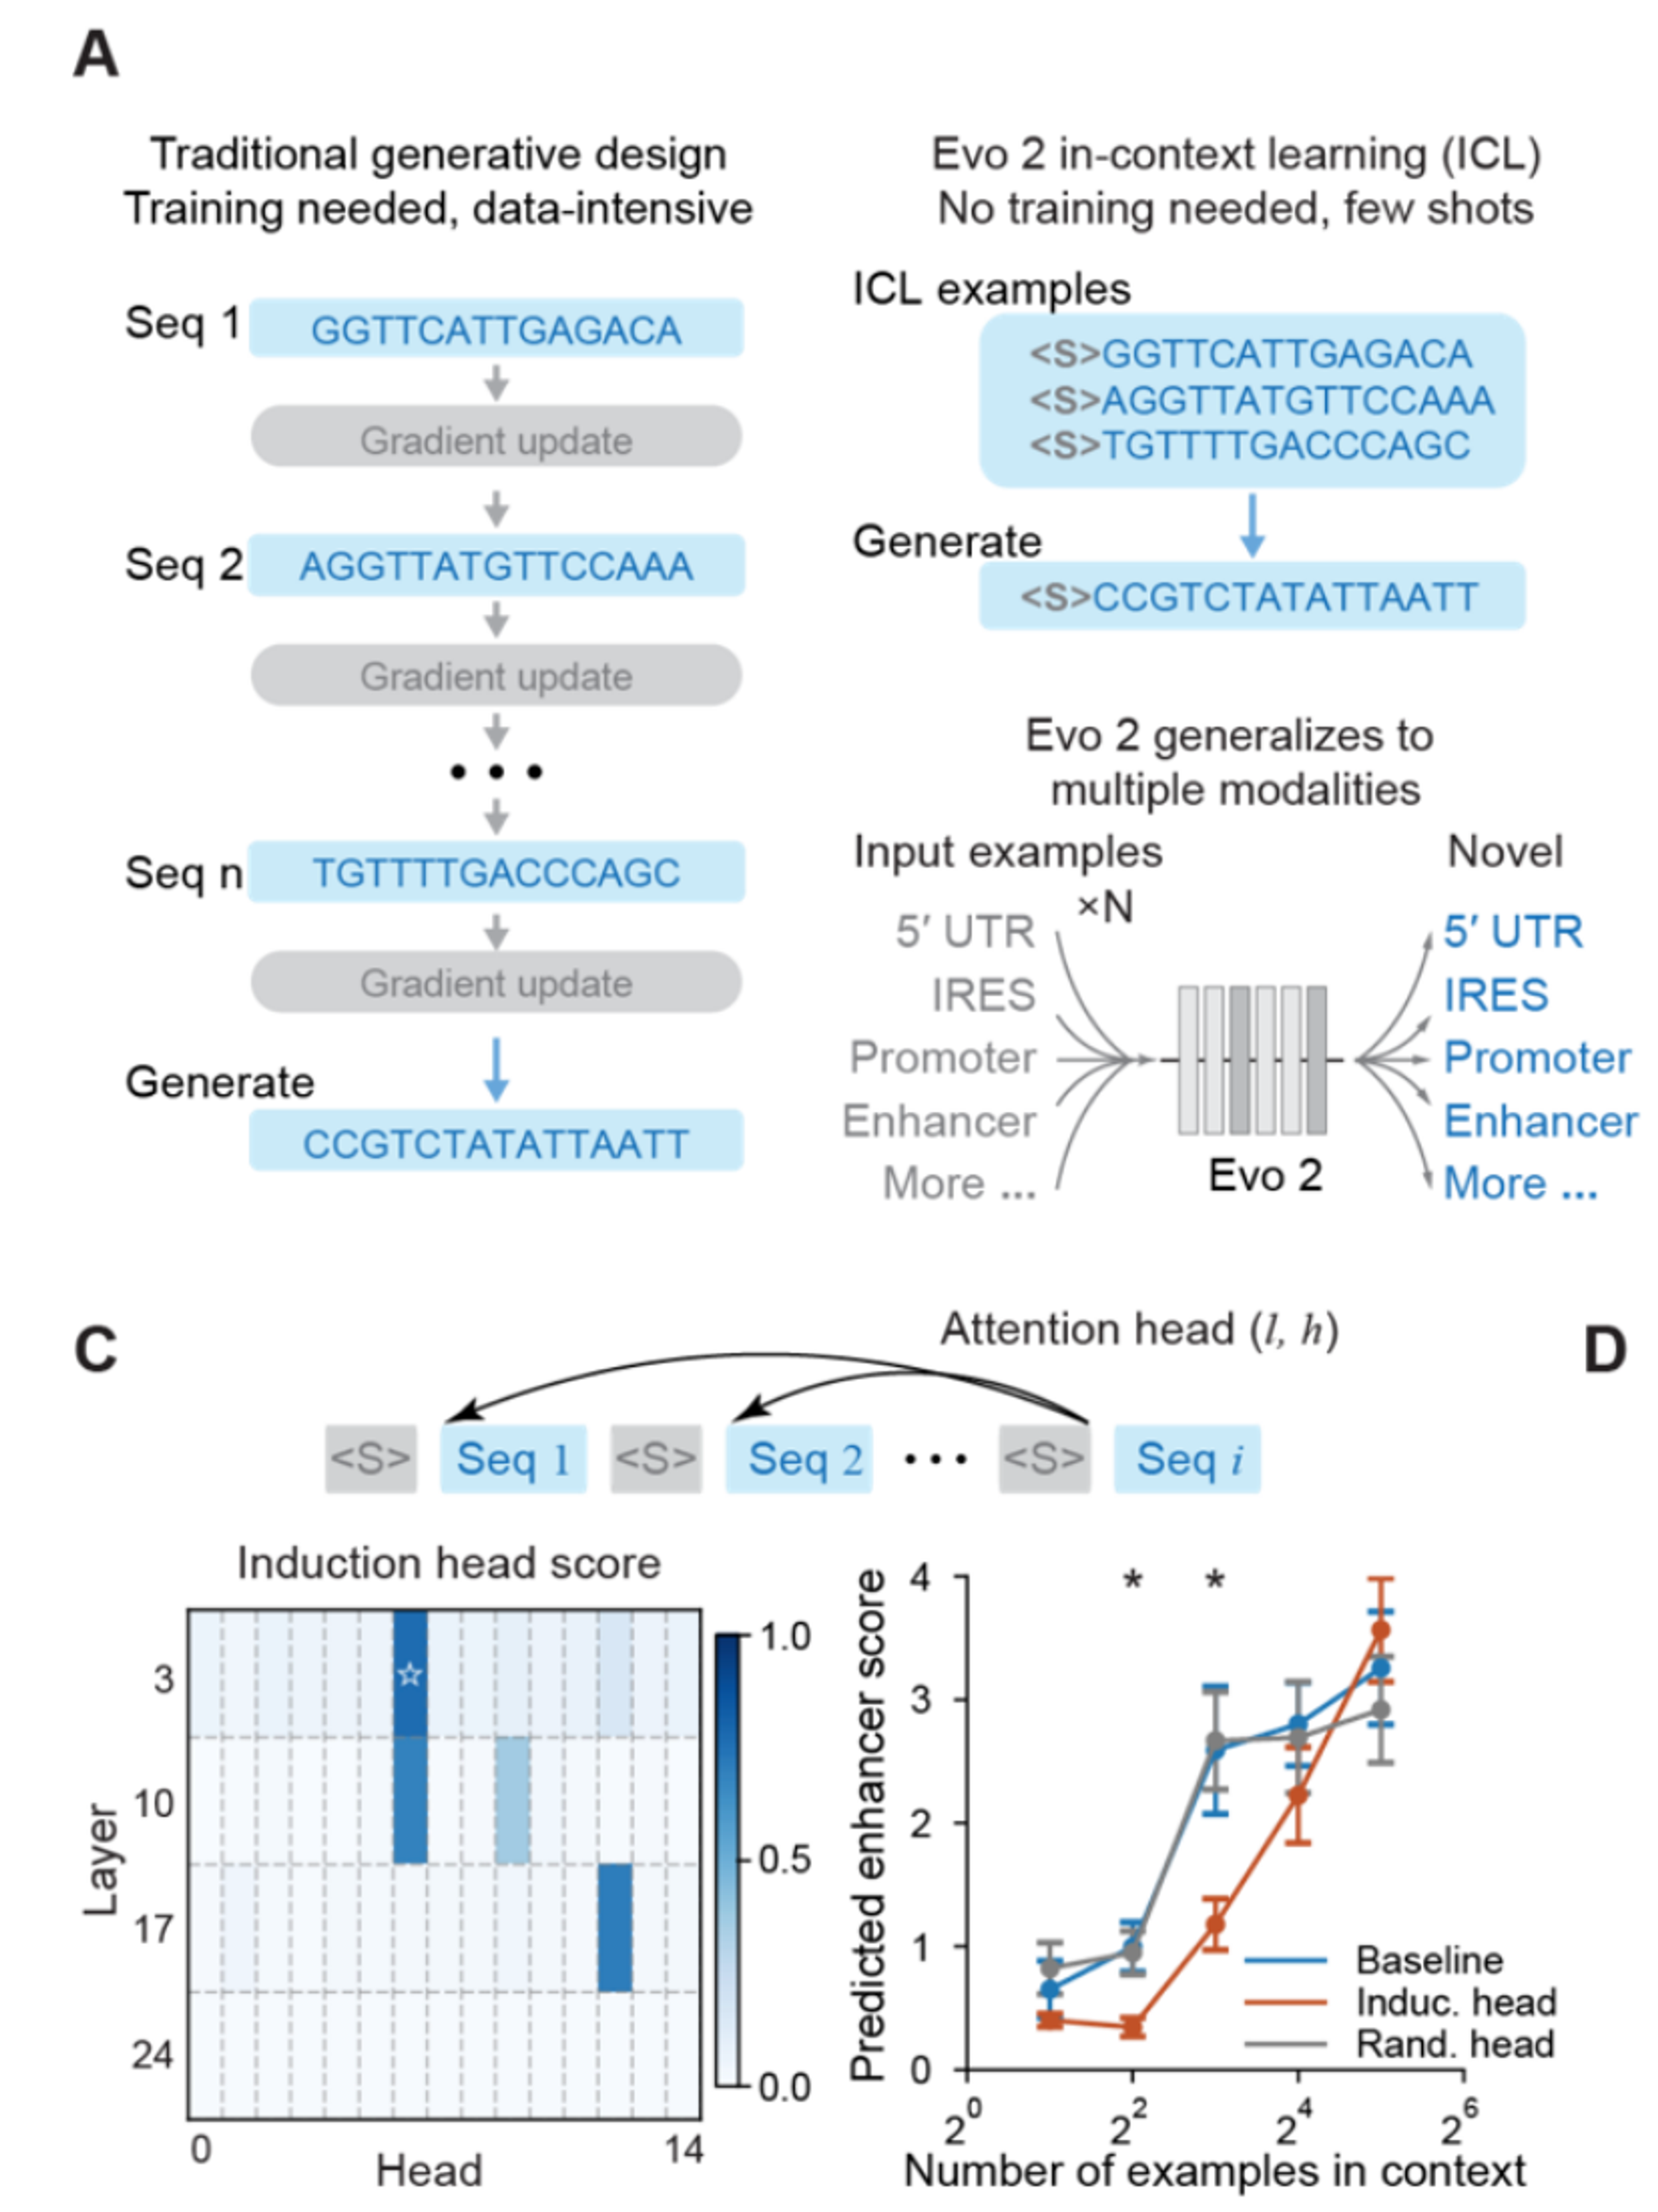
In-context learning (ICL) is an emergent capability of large language models that allows them to adapt to new tasks from a few examples without additional training, enabling these models to become general-purpose learners. Here, we show that the genome language model, Evo 2, exhibits generative ICL in biological sequence design.
Efficient biological design via genomic in-context learning
BioRxiv
Hie Lab

Efficient biological design via genomic in-context learning

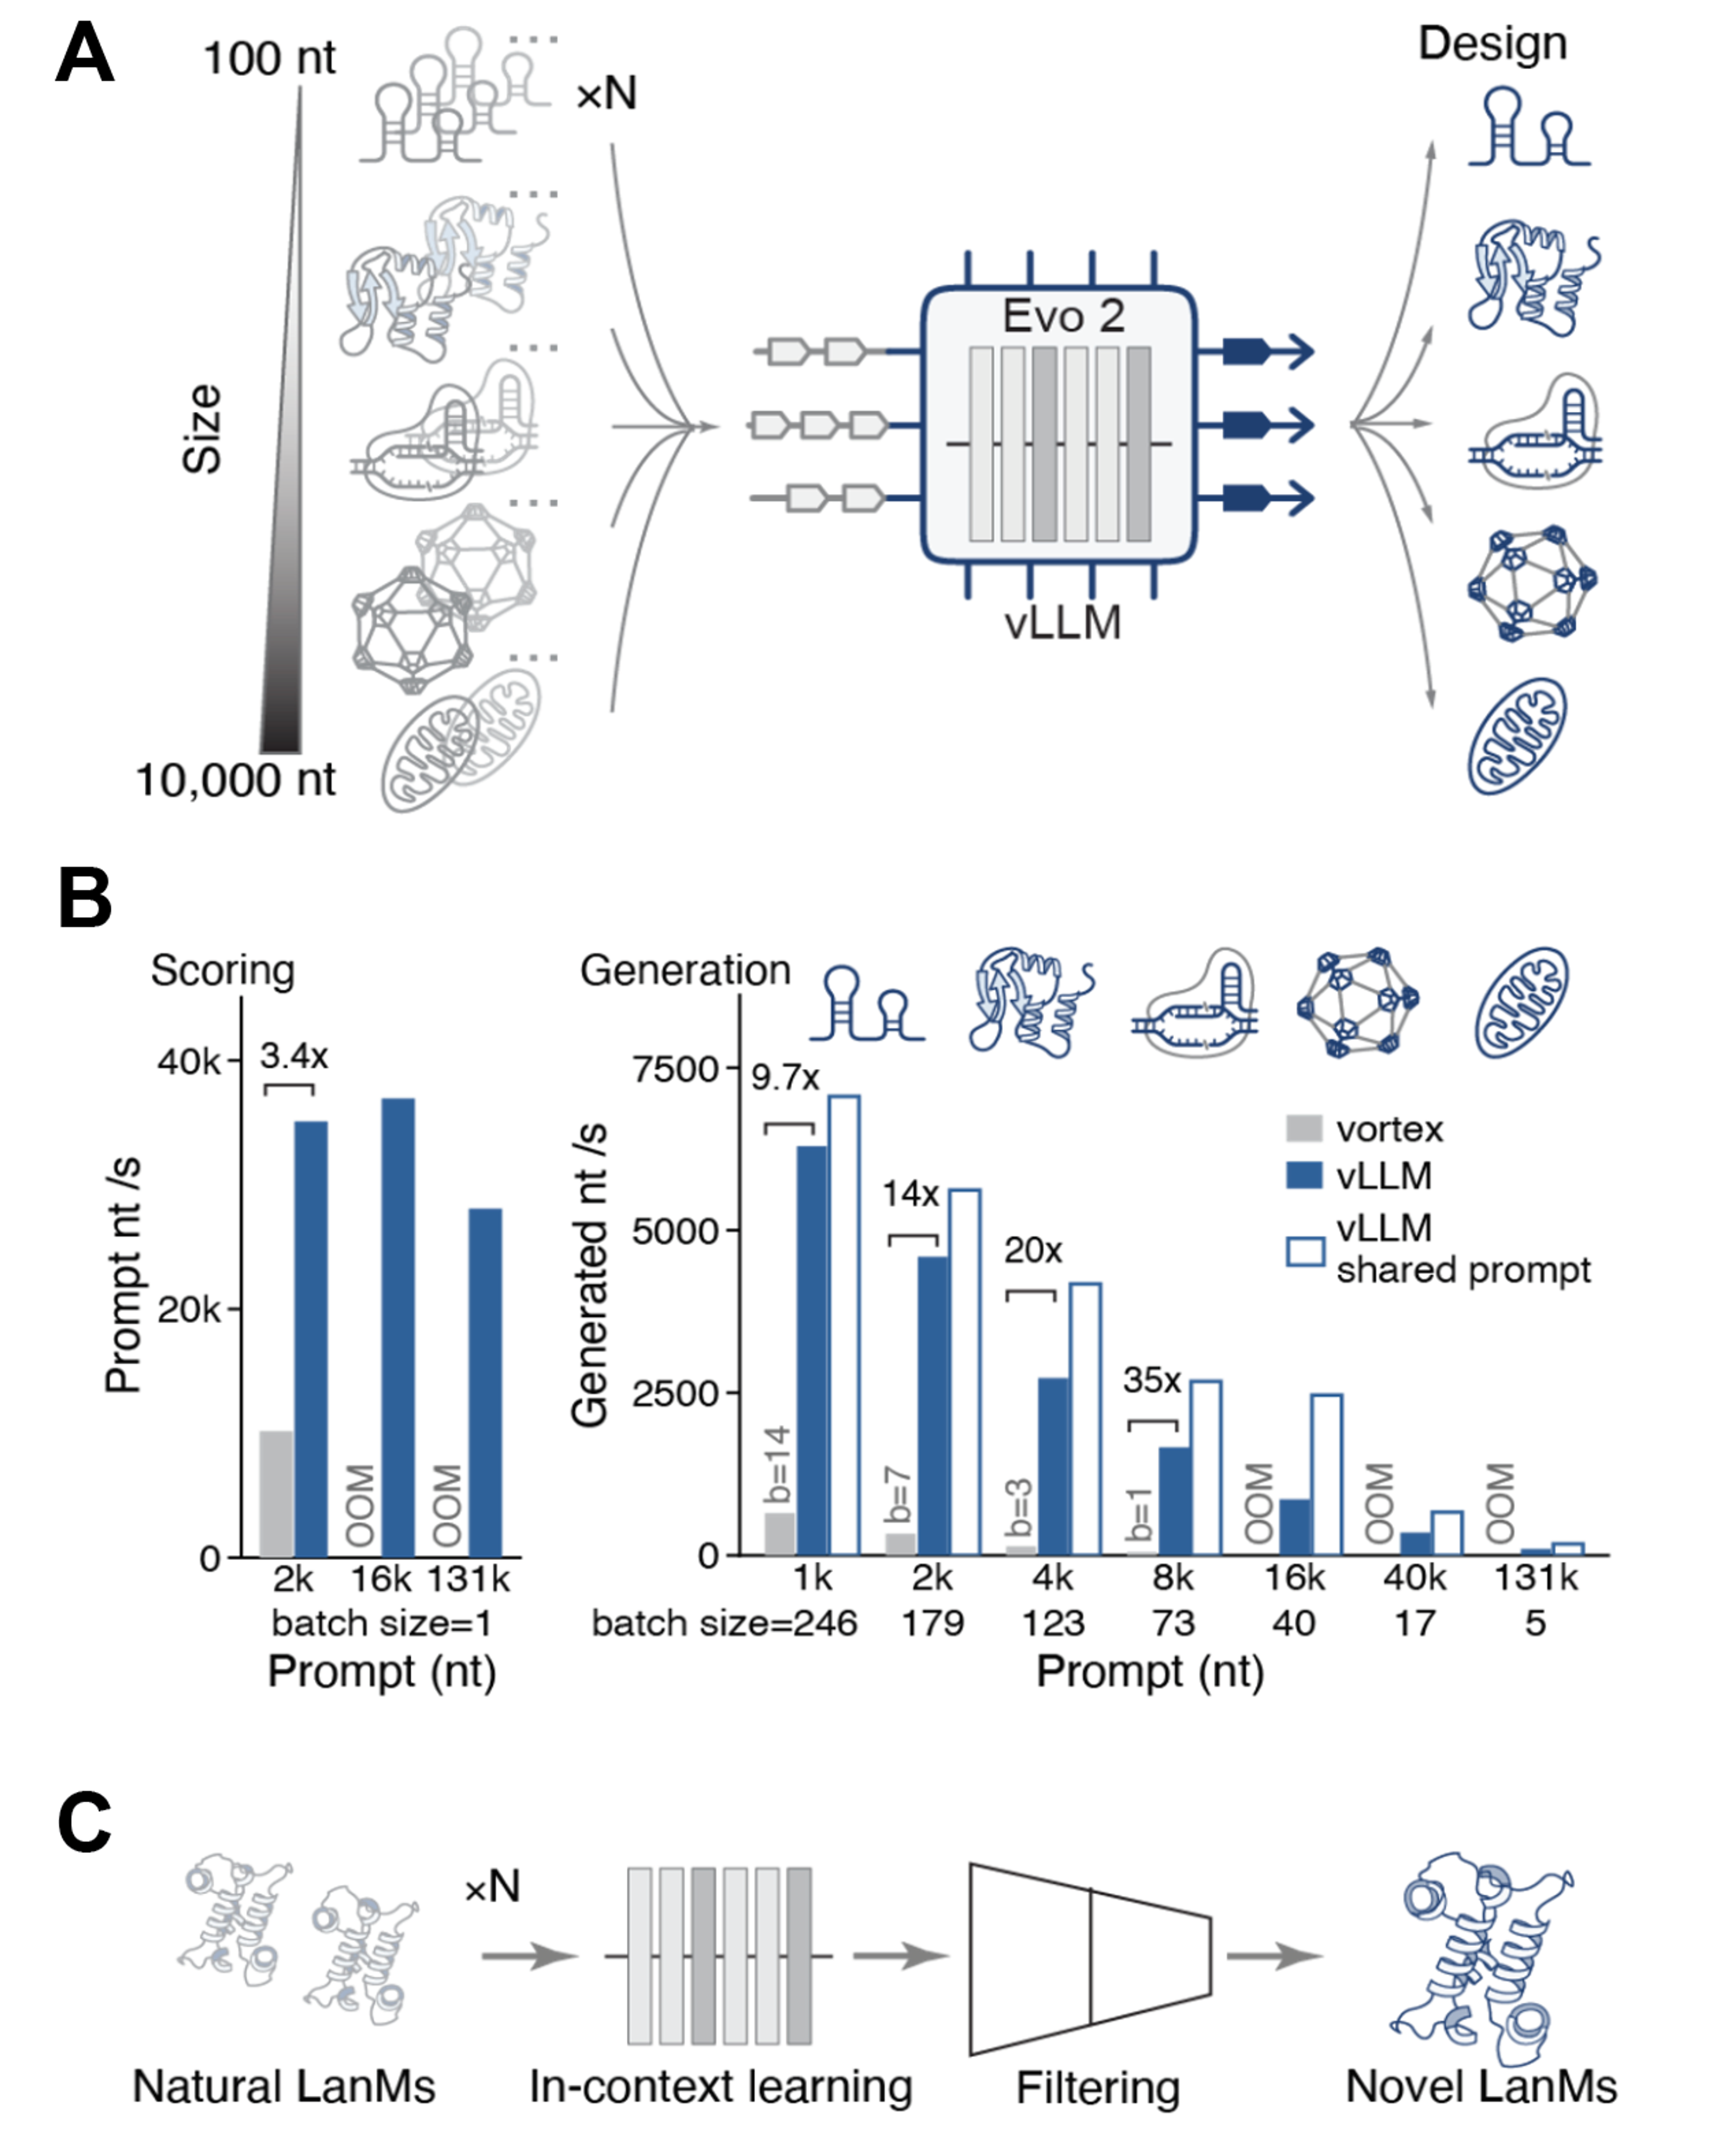
In-context learning (ICL), the ability to perform new tasks from a few examples supplied in a prompt, is a defining emergent capability of large language models. Here, we show that ICL emerges in the genome language model Evo 2 and enables biological design.

Systematic mapping of the prokaryotic noncoding sequence space with a genomic language model
BioRxiv
Hsu LabKonermann Lab

Systematic mapping of the prokaryotic noncoding sequence space with a genomic language model

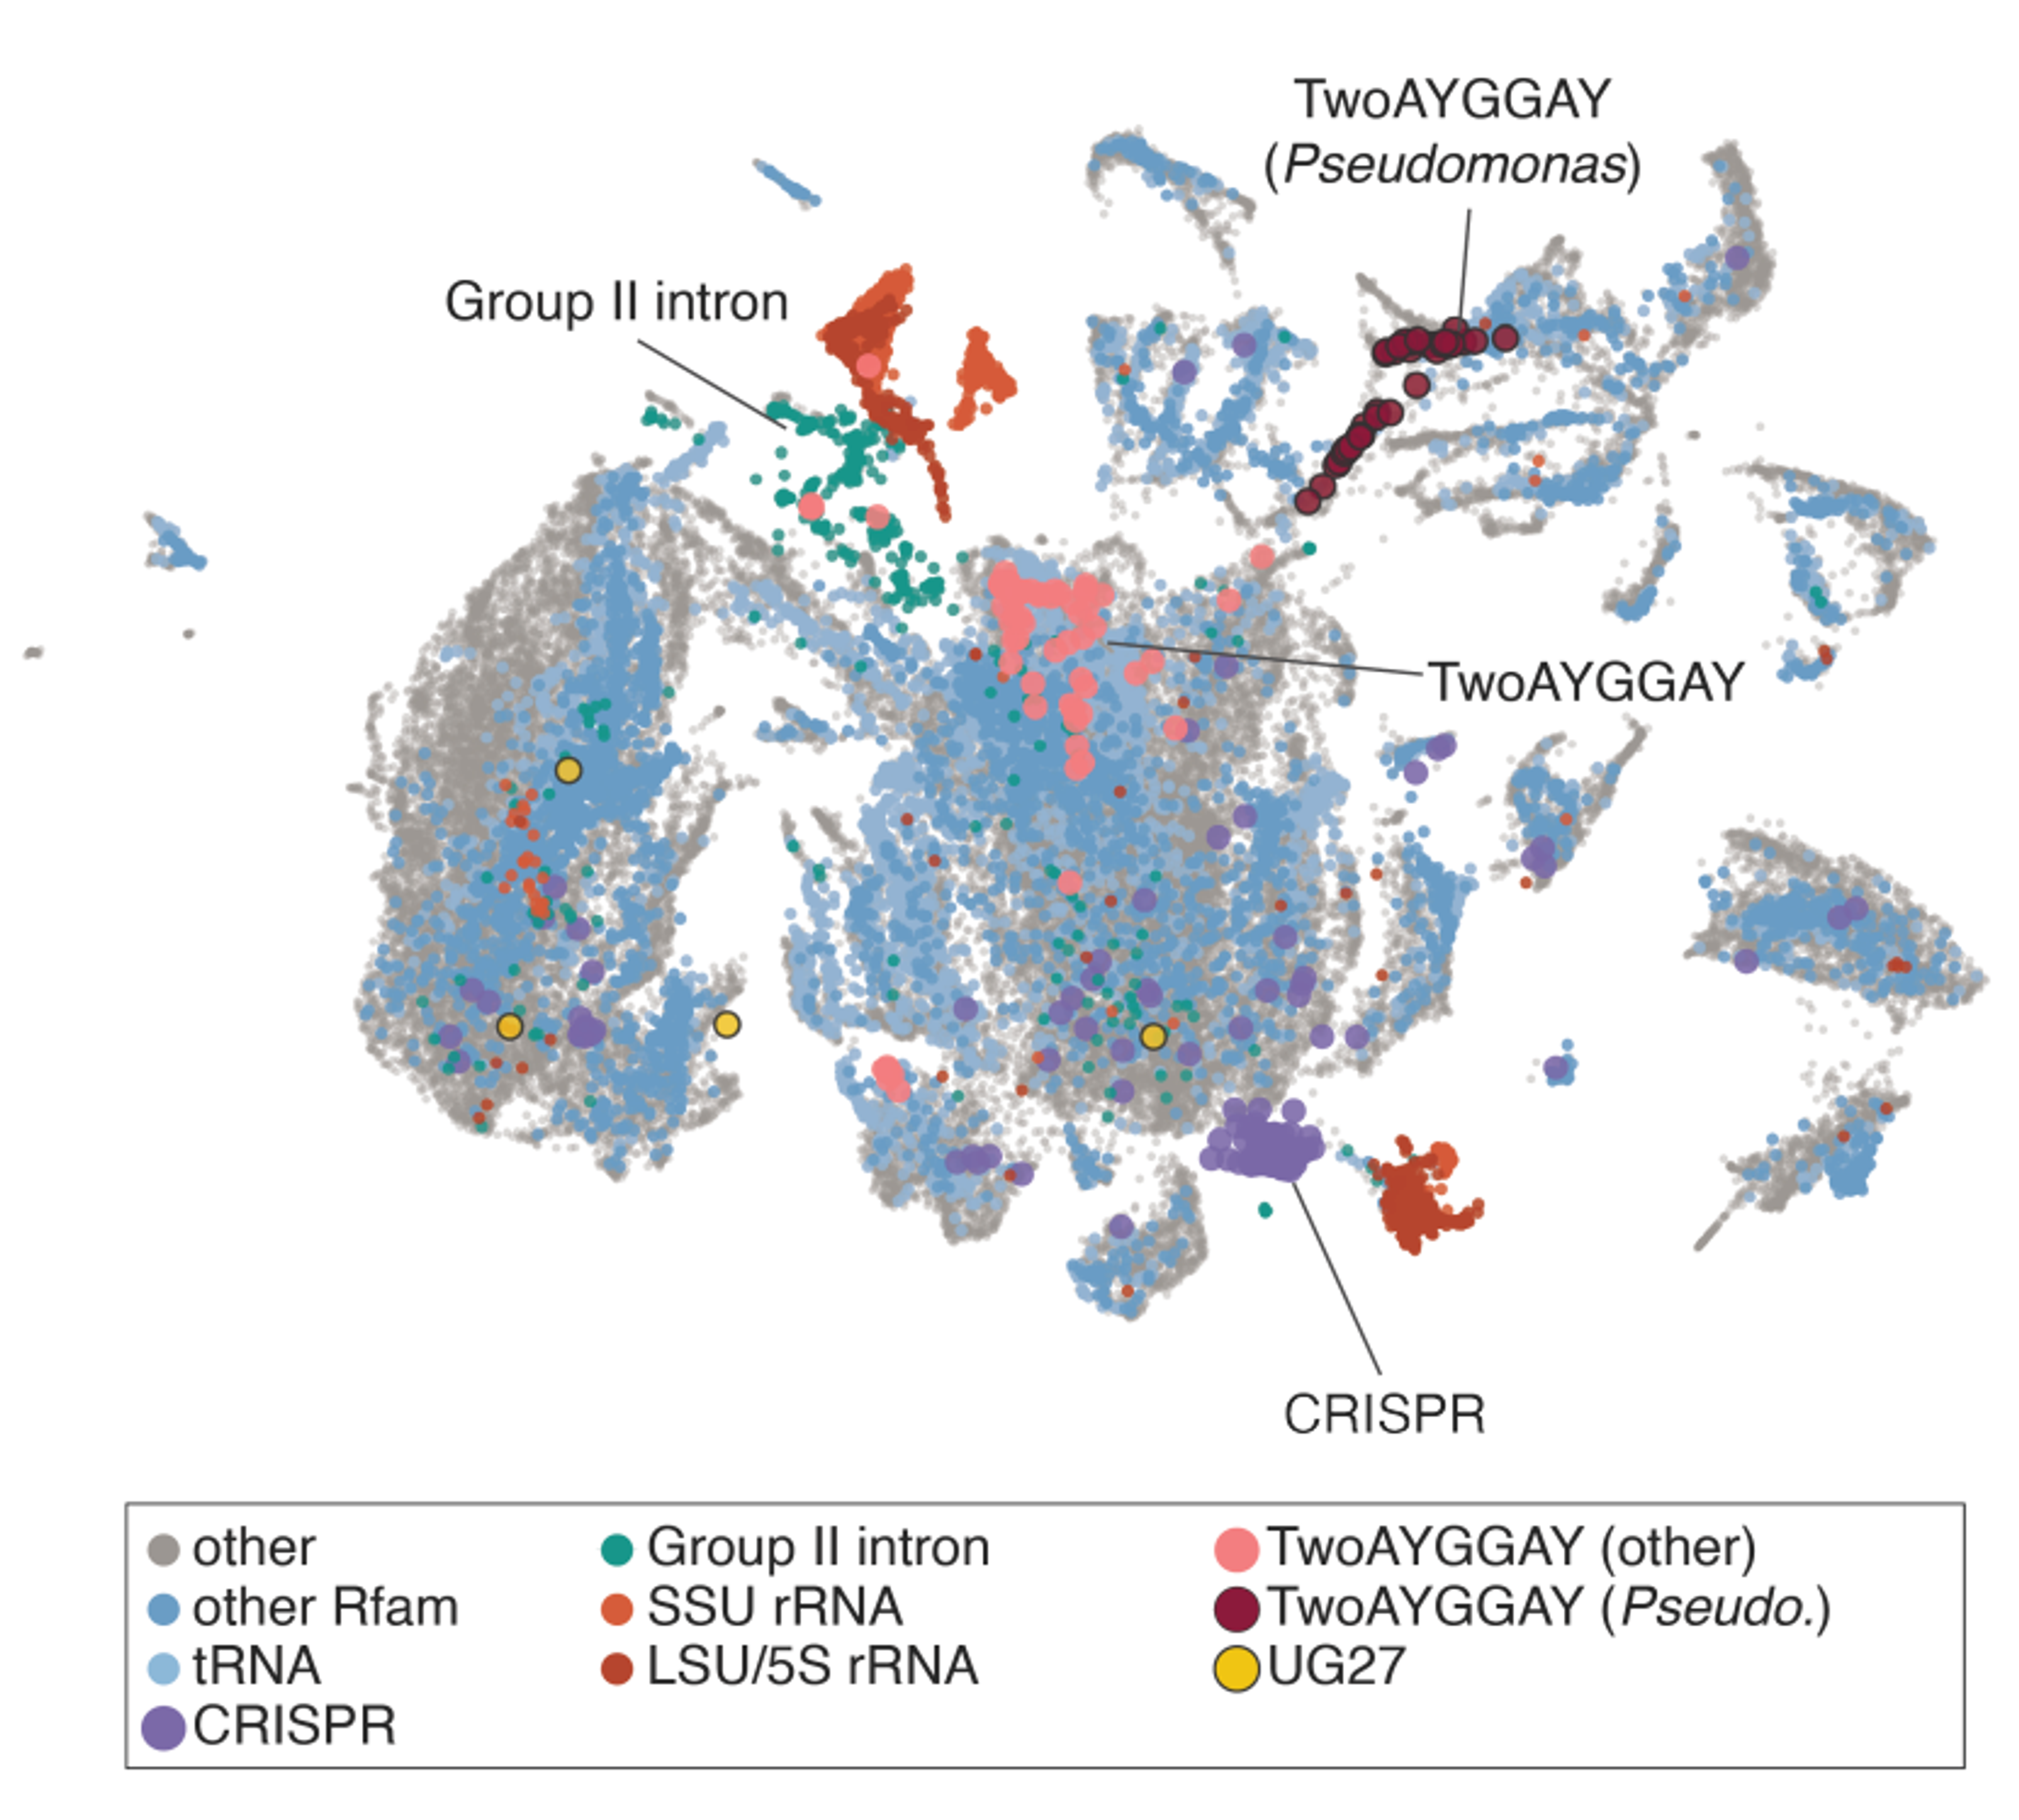
Prokaryotic genomes encode a vast repertoire of noncoding elements such as ribozymes, riboswitches, and CRISPR RNAs. Here, we show that a genomic language model fine-tuned on prokaryotic intergenic sequences learns representations of noncoding elements from DNA sequence.

Predicting cell-state-specific gene regulation from DNA across species
BioRxiv
Zhou Lab

Predicting cell-state-specific gene regulation from DNA across species

We developed Bolero, a sequence-to-function model whose parameters condition on atlas-scale cell-state embeddings and additional biological variables to predict cell-state-specific chromatin accessibility and transcript abundance from DNA.

A high-resolution protein-RNA binding atlas for structure-aware interaction prediction and post-transcriptional regulatory modeling
BioRxiv
Goodarzi Lab

A high-resolution protein-RNA binding atlas for structure-aware interaction prediction and post-transcriptional regulatory modeling

RNA-binding proteins control post-transcriptional gene expression by regulating RNA processing, localization, stability, and translation. Here we introduce a high-throughput CLIP-seq platform that combines endogenous protein tagging for uniform affinity capture with ligation-free library preparation to generate high-resolution RNA-protein binding profiles.

The adaptive architecture of tRNA dependencies across physiological tumor environments
BioRxiv
Goodarzi Lab

The adaptive architecture of tRNA dependencies across physiological tumor environments

The genetic code is decoded by a highly redundant transfer RNA (tRNA) repertoire. Here, we establish an atlas of tRNA dependencies through large-scale CRISPRi perturbation mapping across matched in vivo tumor growth and in vitro cell culture for 16 cancer cell lines spanning seven tissue types.

PIE: Generalizing perturbation effects across unseen perturbations, contexts and datasets
BioRxiv
Computational

PIE: Generalizing perturbation effects across unseen perturbations, contexts and datasets

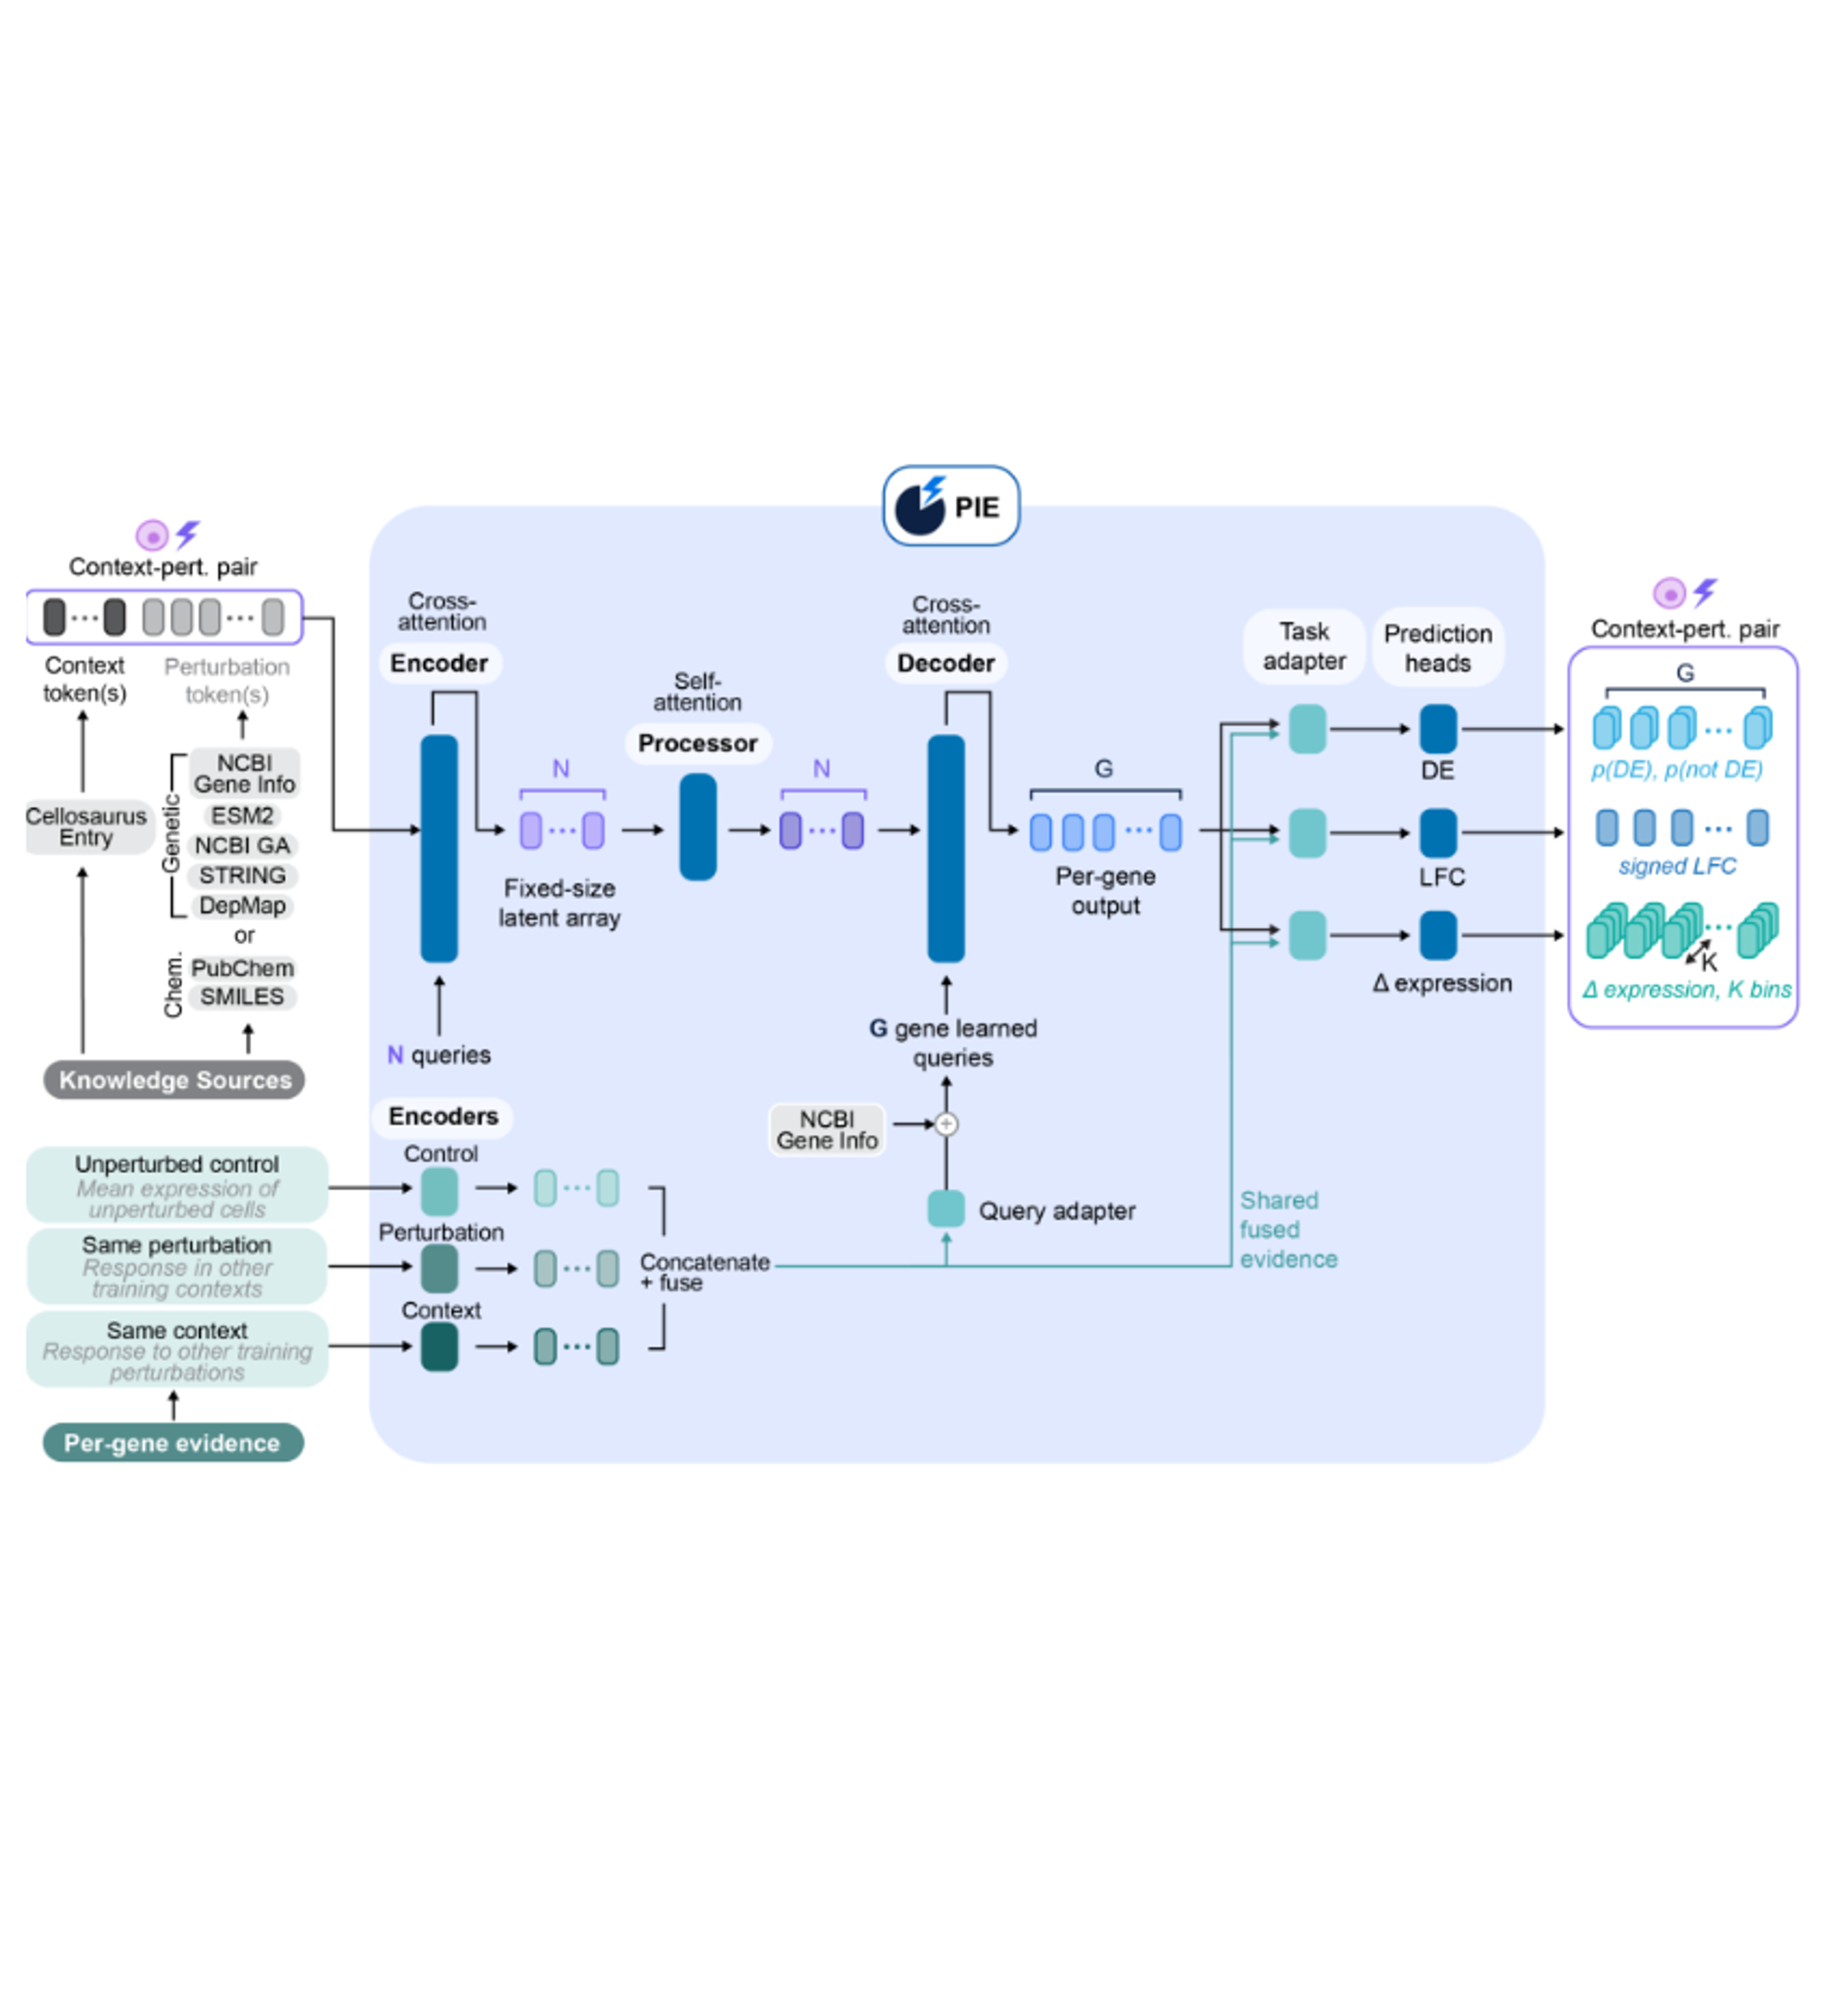
Predicting cellular response to perturbations is key to understanding biological mechanisms and selecting therapeutic targets. Here, we introduce PIE, a model that addresses these limitations by reformulating the learning task around population level perturbation effects and incorporating auxiliary inputs that better characterize the biological system.

Extracellular cGAMP transmission exacerbates neuroinflammation in aging and multiple sclerosis
BioRxiv
Li Lab

Extracellular cGAMP transmission exacerbates neuroinflammation in aging and multiple sclerosis

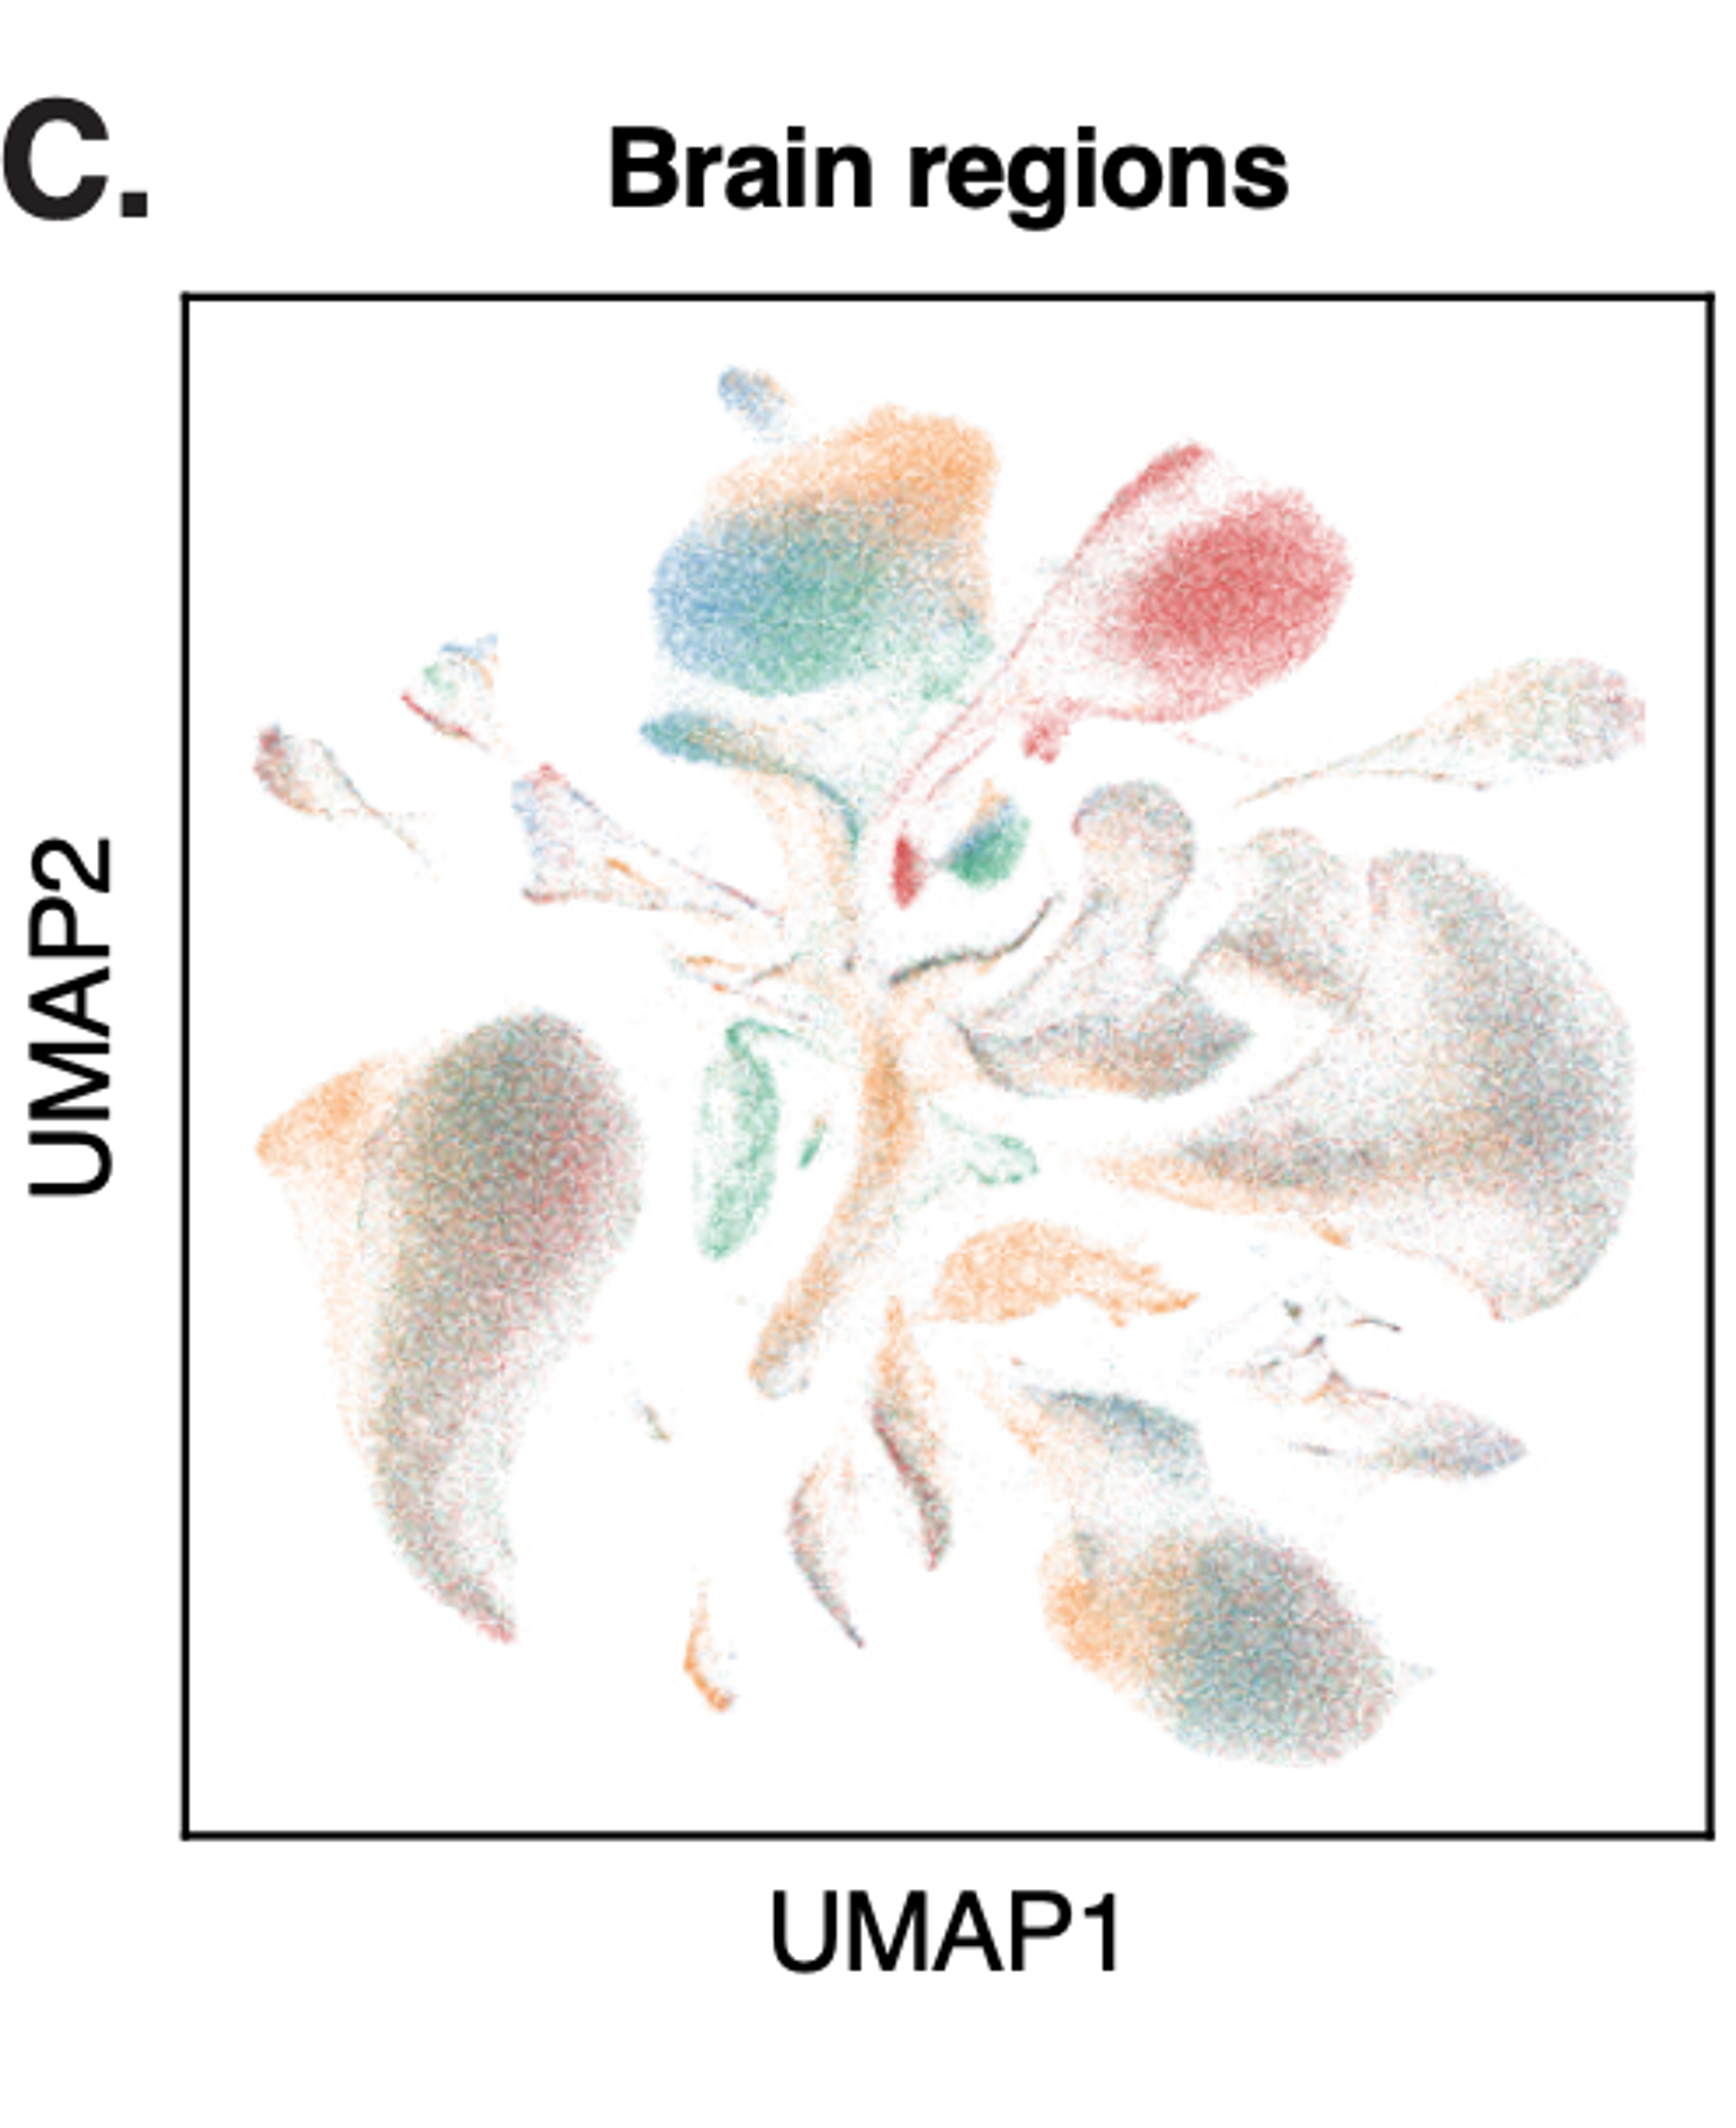
Neuroinflammation drives neurodegeneration and cognitive decline in aging, but the signals that spread this inflammation across the CNS remain unclear. Here, we show in mice that advanced age and experimental autoimmune encephalomyelitis both result in elevated cytosolic mtDNA in microglia.

Coevolutionary mining of prokaryotic non-coding elements with a genome language model
BioRxiv
Hie Lab

Coevolutionary mining of prokaryotic non-coding elements with a genome language model

Microbial genomes encode essential ncRNAs and nucleic-acid-interacting proteins. However, discovery of these systems remains based on similarity to known sequences. Here, we introduce Minerva, a framework for coevolutionary mining that uses genome language models to predict pairwise interactions directly from sequence.

Tahoe-100M: Mapping drug-induced molecular phenotypes at single-cell resolution
Cell
Goodarzi LabComputational

Tahoe-100M: Mapping drug-induced molecular phenotypes at single-cell resolution

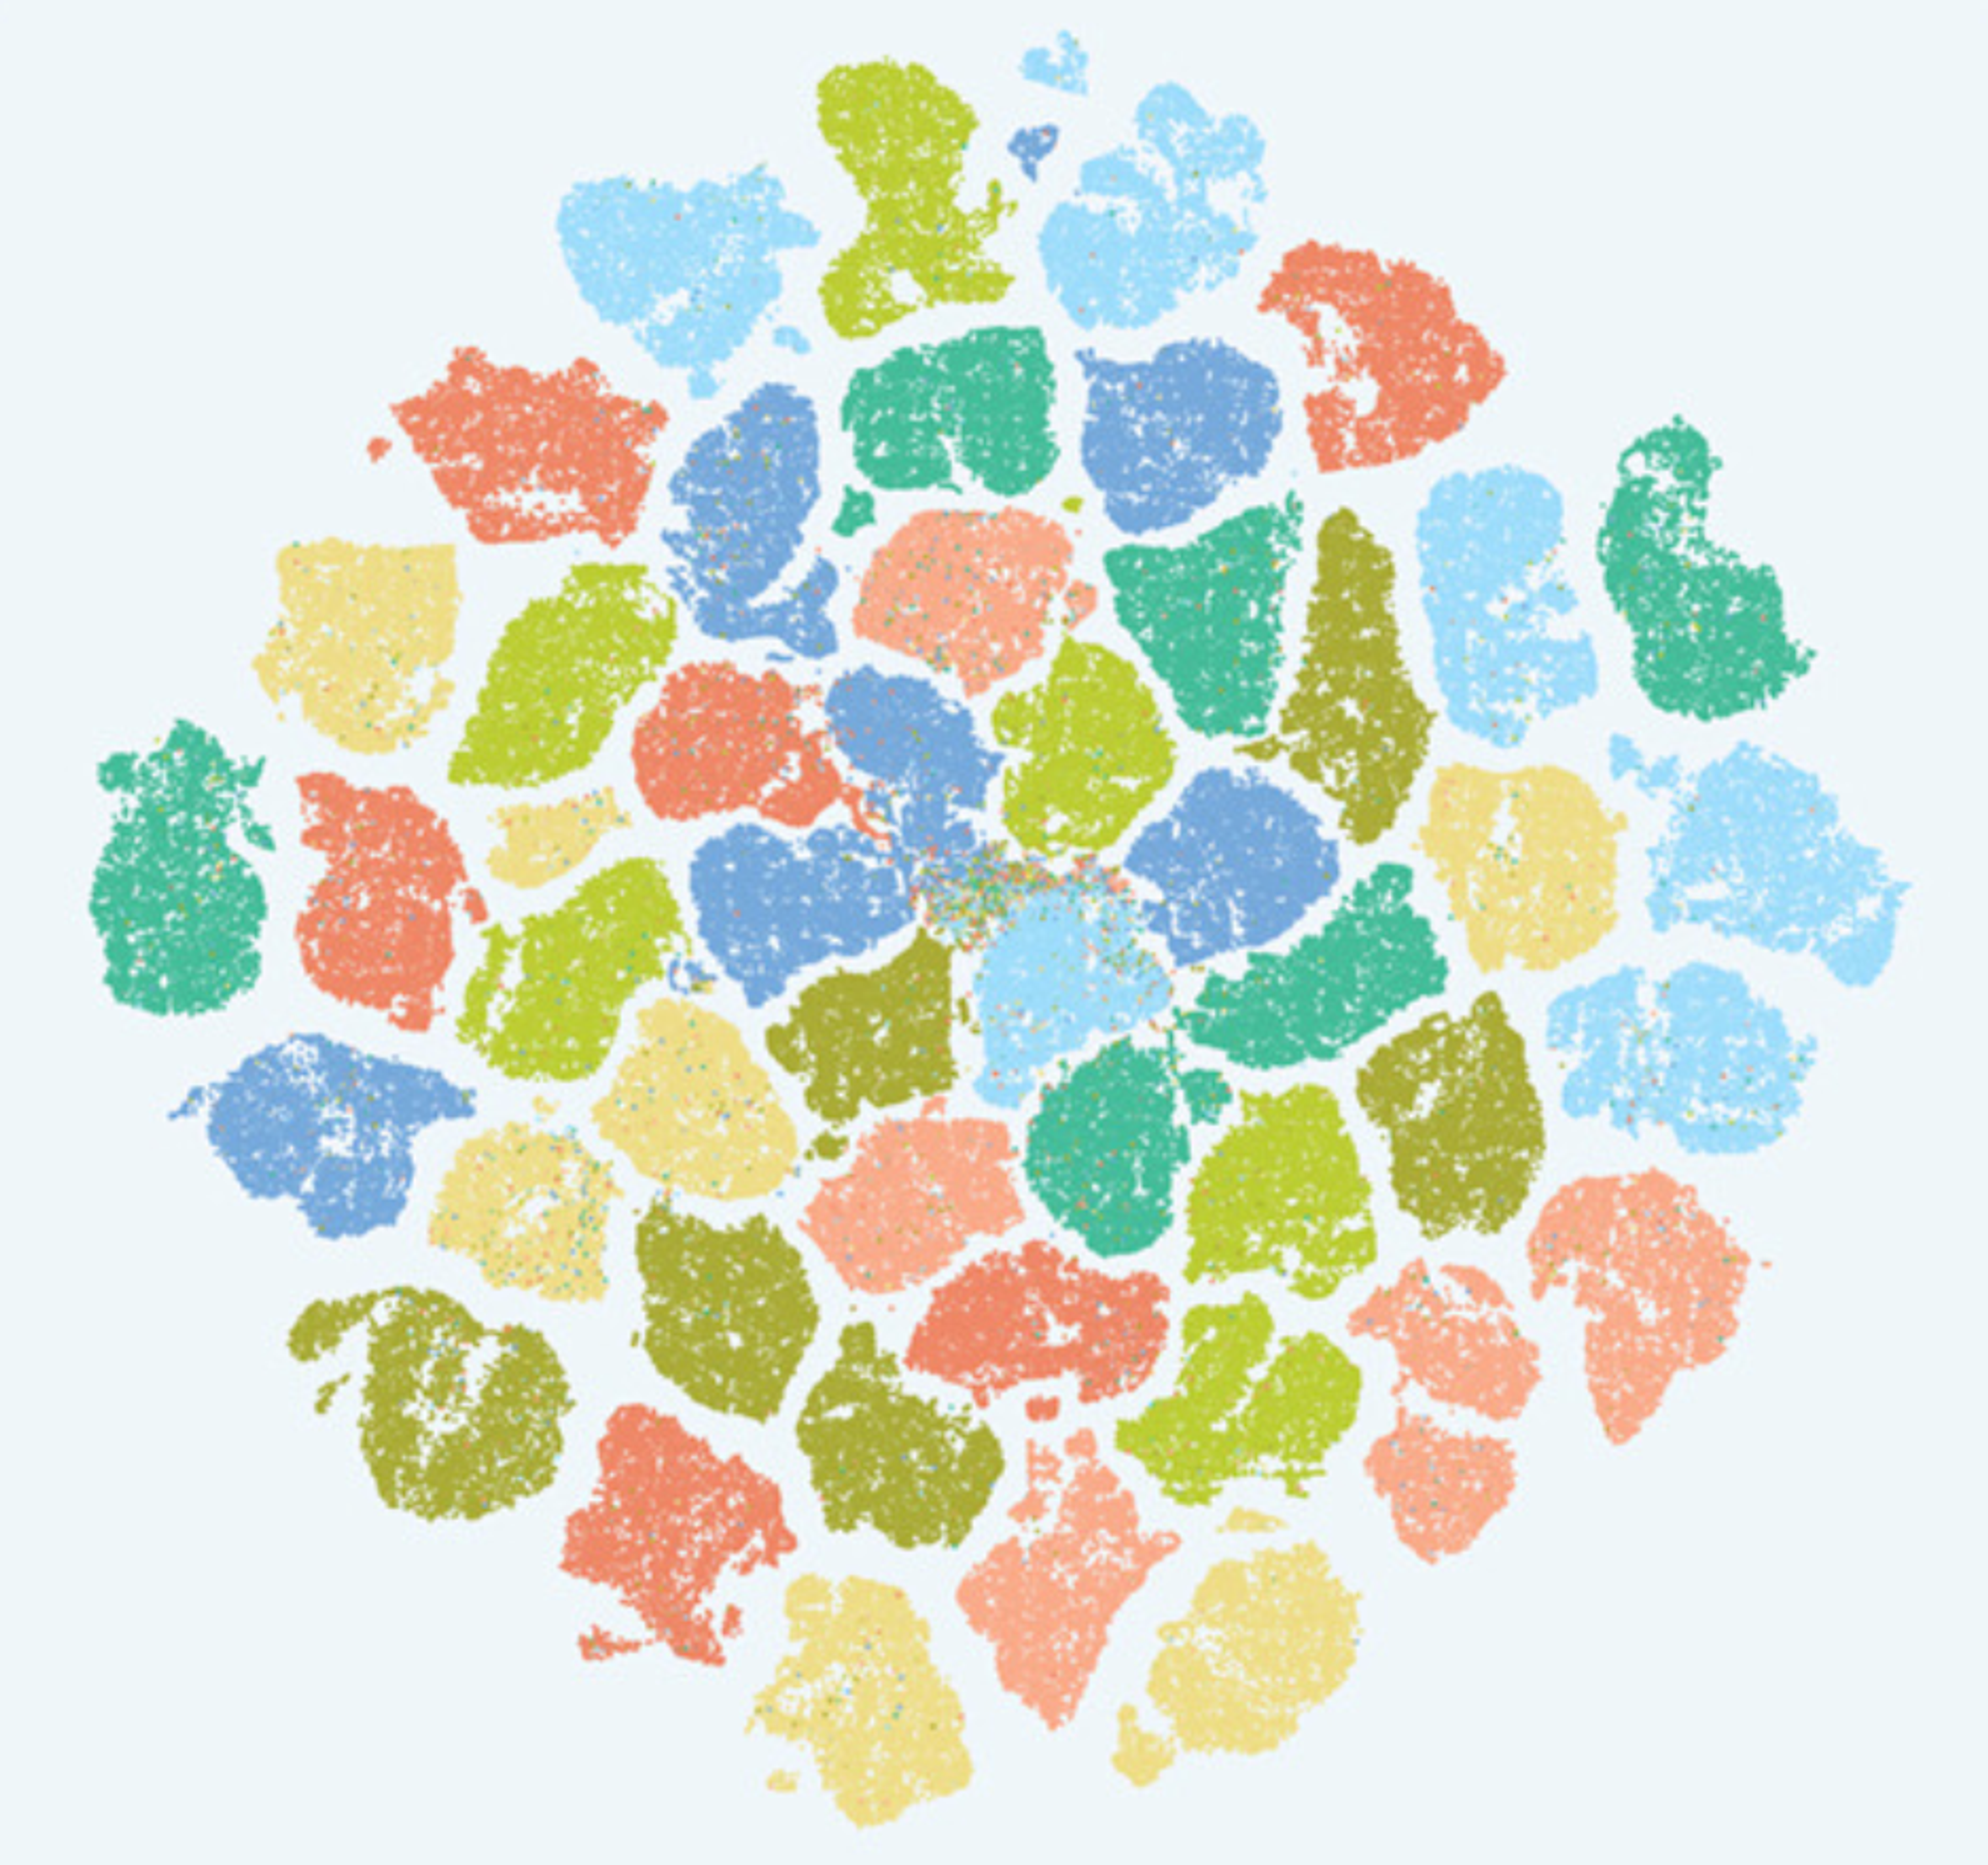
Tahoe-100M, generated by Tahoe, is a giga-scale single-cell perturbation atlas comprising 100 million transcriptomes from 50 diverse cancer cell lines treated with 1,100 drug-dose conditions. This parallel profiling of thousands of perturbations at single-cell resolution with minimal batch effects is enabled by the Mosaic platform.